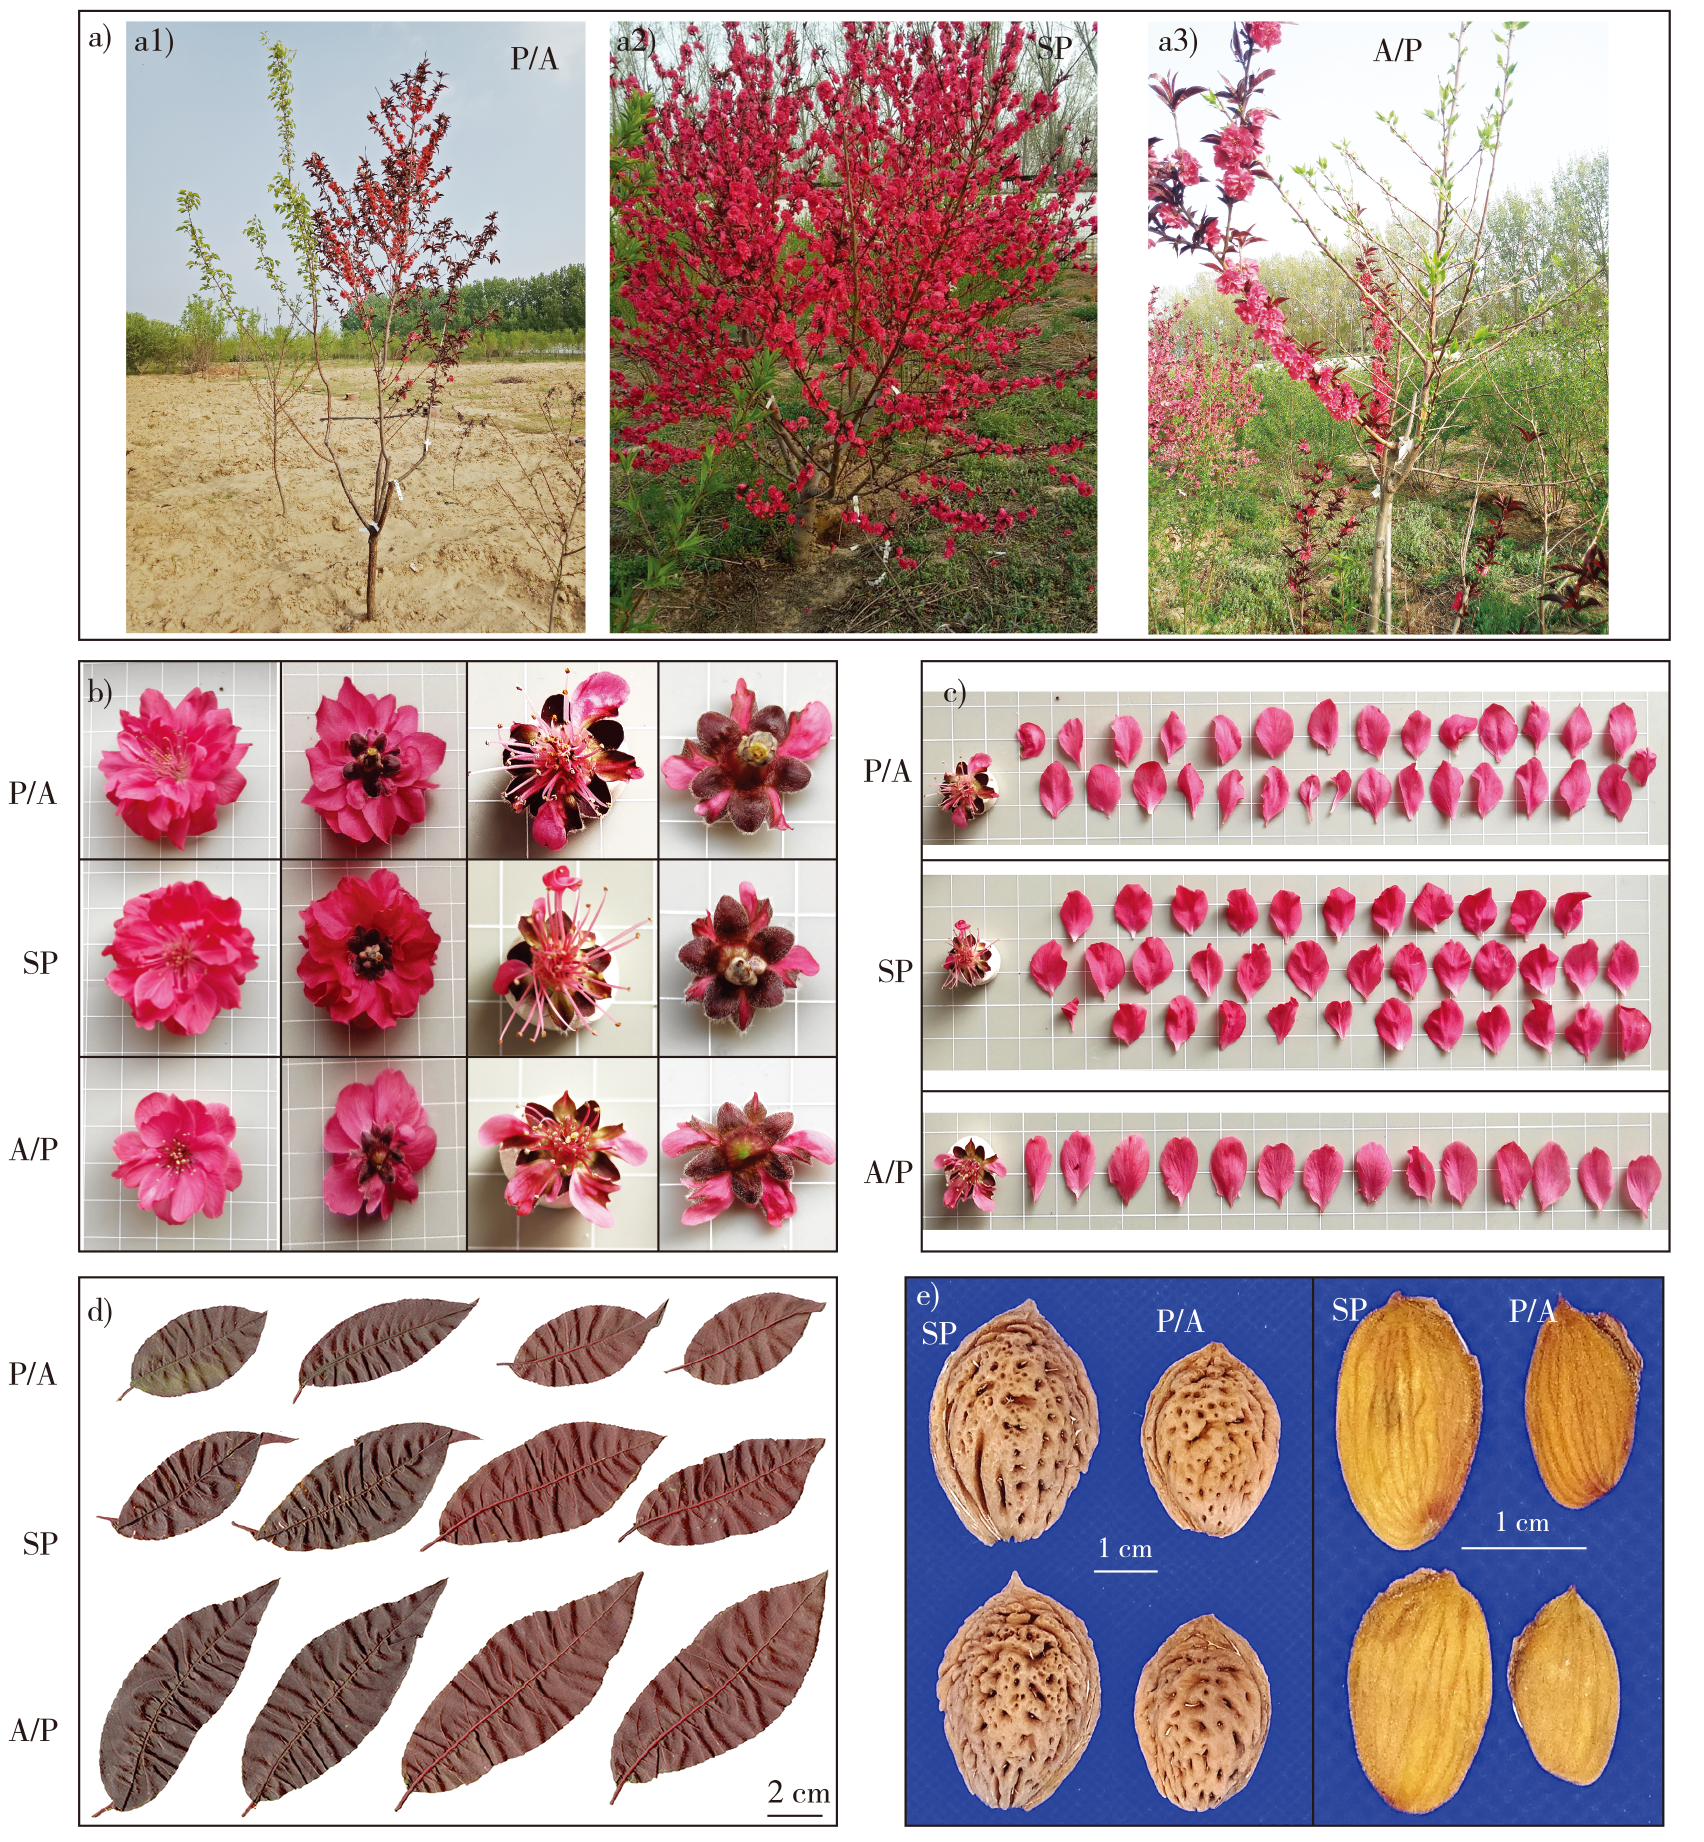
【目的】鉴定影响桃嫁接体差异性状形成的差异表达基因和长链非编码RNA(long non-coding RNA, lncRNA),构建共表达的“lncRNA-mRNA”调控模块,为探索lncRNA在桃、杏嫁接体差异性状形成过程中的作用提供参考。【方法】分别采集桃(Prunus persica)/杏(P. armeniaca)嫁接体(P/A)、杏/桃嫁接体(A/P)和桃本砧嫁接体(SP)中的桃器官(叶、花蕾和完全盛开花)混样后进行RNA-Seq分析,鉴定与嫁接体差异性状相关的差异表达基因(DEGs)和lncRNAs,并进行GO和KEGG功能富集分析,利用Cytoscape构建差异“lncRNA-mRNA”对的共表达网络,并利用qRT-PCR技术对10个差异mRNAs和8个lncRNAs的表达量进行分析。【结果】①与SP相比,从A/P和P/A中分别鉴定到1 115和624个差异基因。其中,特异表达的差异基因分别为942和451个,主要富集在细胞核、质膜、膜的整体成分、叶绿体和细胞质等功能分区。②在A/P vs SP中,富集在植物病原体互作通路上的差异基因最多;而在P/A vs SP中,富集在戊糖和葡萄糖醛酸的相互转化通路上的差异基因最多。③在A/P和P/A中分别有29和26个差异表达的lncRNAs,其中在A/P和P/A特异表达的分别为21和18个。④构建24个差异表达的“lncRNA-mRNA”对,其中A/P和P/A中分别有14和10个。⑤通过基因的功能注释发现,A/P中的MSTRG.17020.2-XM_007210198.2可能参与氨酰的生物合成,MSTRG.8395.1-XM_007217967.2可能参与半乳糖代谢,P/A中的MSTRG.6365.3-XM_020556240.1/XM_020556234.1可能参与嫁接体的防御反应过程。【结论】从桃嫁接体中鉴定出与其差异性状相关的差异mRNAs和lncRNAs,并发现关键的共表达“lncRNA-mRNA”调控模块,为进一步研究lncRNAs调控嫁接体差异性状的功能作用奠定基础。
【Objective】This study aims to identify differentially expressed genes (DEGs) and long non-coding RNAs (lncRNAs) associated with phenotypic variation in peach grafting hybrids, and to construct ‘lncRNA-mRNA’ co-expression regulatory modules, thereby providing a theoretical foundation for understanding the role of lncRNAs in the formation of differential traits in peach-apricot grafting systems.【Method】The mixed tissue samples of peach leaves, flower buds, and fully bloomed flowers were collected from three grafting combinations: Prunus persica (peach)/P. armeniaca (apricot) grafted hybrid (P/A), apricot/peach grafted hybrid (A/P), and self-rooted peach (SP). Transcriptome sequencing was performed to identify DEGs and lncRNAs related to grafting-induced phenotypic variation. Gene ontology (GO) and Kyoto encyclopedia of genes and genomes (KEGG) enrichment analyses were conducted to elucidate their biological functions. Co-expression networks of ‘lncRNA-mRNA’ pairs were constructed using Cytoscape software, and the expression levels of ten DEGs and eight lncRNAs were validated by quantitative real-time PCR (qRT-PCR).【Result】Analysis of transcriptome sequencing data revealed that 1 115 DEGs were identified in A/P vs SP, and there are 734 upregulated and 381 downregulated genes in A/P compared to SP, respectively. In P/A vs SP, 624 DEGs were detected, with 415 upregulated and 209 downregulated genes in P/A compared to SP, respectively. Venn analysis revealed that a total of 173 common DEGs between the two comparisons, while 942 and 451 DEGs were specifically expressed in A/P and P/A, respectively. GO enrichment analysis showed that the DEGs were significantly enriched in 25 biological processes (BPs), 15 cellular components (CCs), and 10 molecular functions (MFs). In A/P vs SP, 833 DEGs were concentrated in the functional regions of nucleus, plasma membrane, integral component of membrane, chloroplast and cytoplasm, with the highest number of DEGs likely participating in nuclear regulation. A total of 401 DEGs were potentially involved in BPs, including biological process, regulation of DNA-templated transcription, DNA-templated transcription, oxidation-reduction process, and defense response. Regarding MFs, the DEGs were predominantly enriched in functional categories such as molecular function, protein binding, and D binding transcription factor activity, accounting for 52.37% of the total. Other significant MFs categories included metal ion binding, ATP binding, and sequence-specific D binding. In P/A vs SP, 537 DEGs were potentially associated with CCs, including nucleus, plasma membrane, integral component of membrane, cytoplasm, extracellular region, chloroplast, and cytosol, with the highest number of DEGs participating in nuclear regulation. For BPs, the DEGs mainly participated in biological process, regulation of DNA-templated transcription, DNA-templated transcription, defense response, and oxidation-reduction processes. In MFs, DEGs were primarily involved in molecular function, protein binding, and ATP binding, collectively representing over 50% of the functional distribution. KEGG analysis indicated that DEGs in A/P were primarily involved in plant-pathogen interaction, plant hormone signal transduction, starch and sucrose metabolism, MAPK signaling pathway-plant, and phenylpropanoid biosynthesis. In contrast, DEGs in P/A were mainly enriched in pentose and glucuronate interconversions, phenylpropanoid biosynthesis, and starch and sucrose metabolism. A total of 29 and 26 differentially expressed lncRNAs (DElncRNAs) were identified in A/P vs SP and P/A vs SP, respectively, including 16 upregulated and 13 downregulated lncRNAs in A/P vs SP, and 17 upregulated and 9 downregulated lncRNAs in P/A vs SP. Venn analysis showed that there are 8 common DElncRNAs between the two comparisons, with 21 and 18 DElncRNAs specifically expressed in A/P and P/A, respectively. A total of 24 lncRNA-mRNA pairs were identified, including 14 lncRNA-mRNA pairs in A/P vs SP and 10 lncRNA-mRNA pairs in P/A vs SP, with 13 positive and 11 negative regulatory relationships. For example, MSTRG.18445.1 was downregulated, while its target gene XM_007207084.2 was upregulated in A/P compared to SP. MSTRG.17020.2 was downregulated in A/P compared to SP, XM_007210198.2 and XM_00721476.2 were upregulated in A/P compared to MSTRG.8395.1 and XM_007217967.2 both were upregulated in A/P compared to SP. In P/A vs SP, MSTRG.6365.3 was downregulated, while its targets XM_020556240.1 and XM_020556234.1 were upregulated. Based on functional annotation of the DEmRNAs, it was revealed that MSTRG.17020.2 might participate in aminoacyl biosynthesis by negatively regulating the expression level of XM_007210198.2 in A/P, while MSTRG.8395.1 could be involved in galactose metabolism through positive regulation of XM_007217967.2 expression. In P/A, MSTRG.6365.3 appeared to contribute to plant defense responses by negatively regulating the expression of either XM_020556240.1 or XM_020556234.1. qRT-PCR validation of randomly selected 10 DEmRNAs and 8 DElncRNAs from the co-expressed lncRNA-mRNA pairs demonstrated that both the results of RNA-Seq and the constructed lncRNA-mRNA co-expression network were reliable, which provides a solid foundation for further exploration and analysis of key genes involved in the growth and development of apricot-peach grafting hybrids. 【Conclusion】 In this study, some key differentially expressed genes and lncRNAs associated with phenotypic variation in peach grafting hybrids were identified, and key lncRNA-mRNA co-expression regulatory modules were constructed. These findings provide valuable insights into the molecular mechanisms underlying grafting-induced trait formation, and lay a foundation for further research on the functional roles of lncRNAs in fruit tree grafting.
 PDF(55446 KB)